We don't just advise on your market access, we take charge of it
Certified medical device manufacturer: we take charge of your CE marking
Regulatory compliance remains a major challenge for manufacturers. As a medical device manufacturer ourselves, we bring a certified QMS, clinical evaluation, technical documentation and notified body relations. We support you in bringing your product to market.
ISO 13485 certification of the quality management system.
2015
First CE markings obtained on our devices.
2017
Integration of 3D printing of custom-made medical devices.
2019
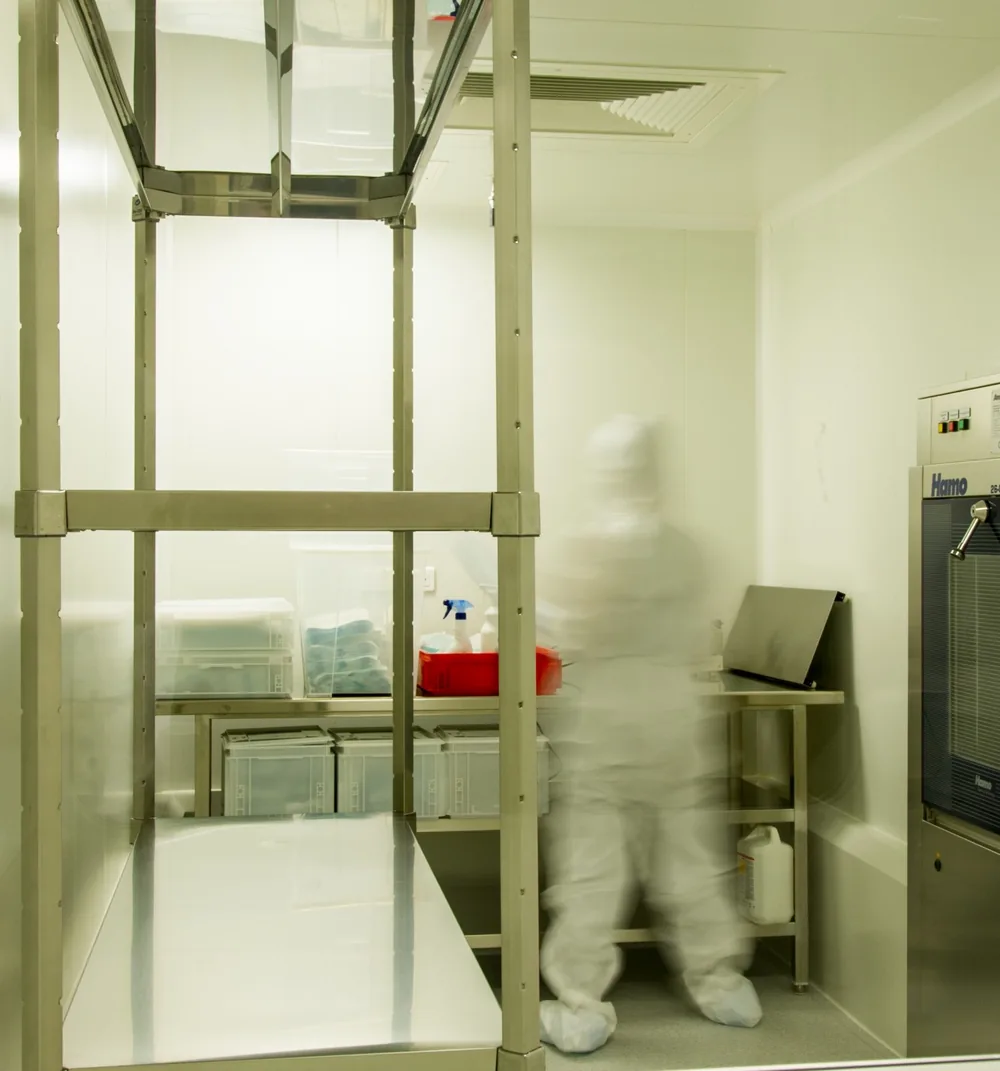
Integration of an ISO 7 cleanroom at our production site.
2025
MDR transition completed for our entire off-the-shelf orthopedic product range.
2026
First 510(k) clearance obtained from the FDA.
Today
Certificates obtained under Regulation (EU) 2017/745 (MDR). Class I, Im and IIa devices, and custom-made devices.
Notified body: GMED, identification number 0459.
Expertise in Class IIb and III devices, gained through bringing several implant ranges to market for our clients.
Scope
What our manufacturer status covers
Four areas, all operated in-house.
01
QMS
ISO 13485-certified QMS since 2014
Processes documented, audited and maintained over time
Risk management and traceability from design to production
Software development carried out according to the requirements applicable to medical device software
02
Technical file and CE marking
Compilation and maintenance of the technical file under MDR 2017/745
Established relationship with our notified body
Clinical evaluation and drafting of the PMCF plan (clinical investigation, user surveys)
Compilation of your CE technical file within our scope, transferable if neededv
03
Additive manufacturing
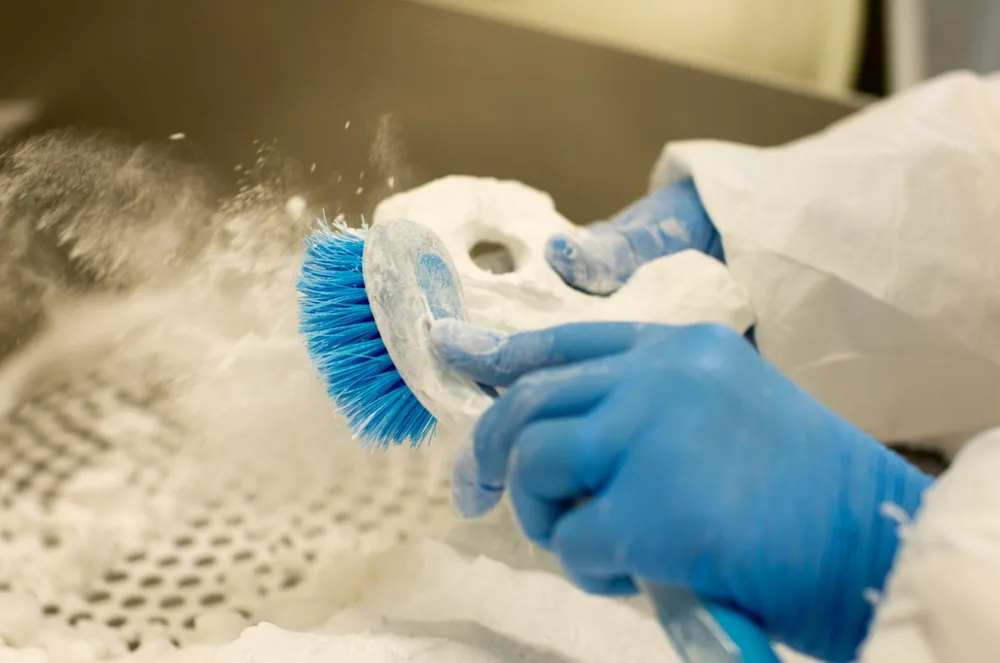
3D printing of custom-made medical devices since 2017
Integrated ISO 7 cleanroom, on our own site
Ready-to-sterilize packaging, or gamma sterilization
Patient-specific intraoperative guides and instruments
04
Post-market
Vigilance
Post-market surveillance
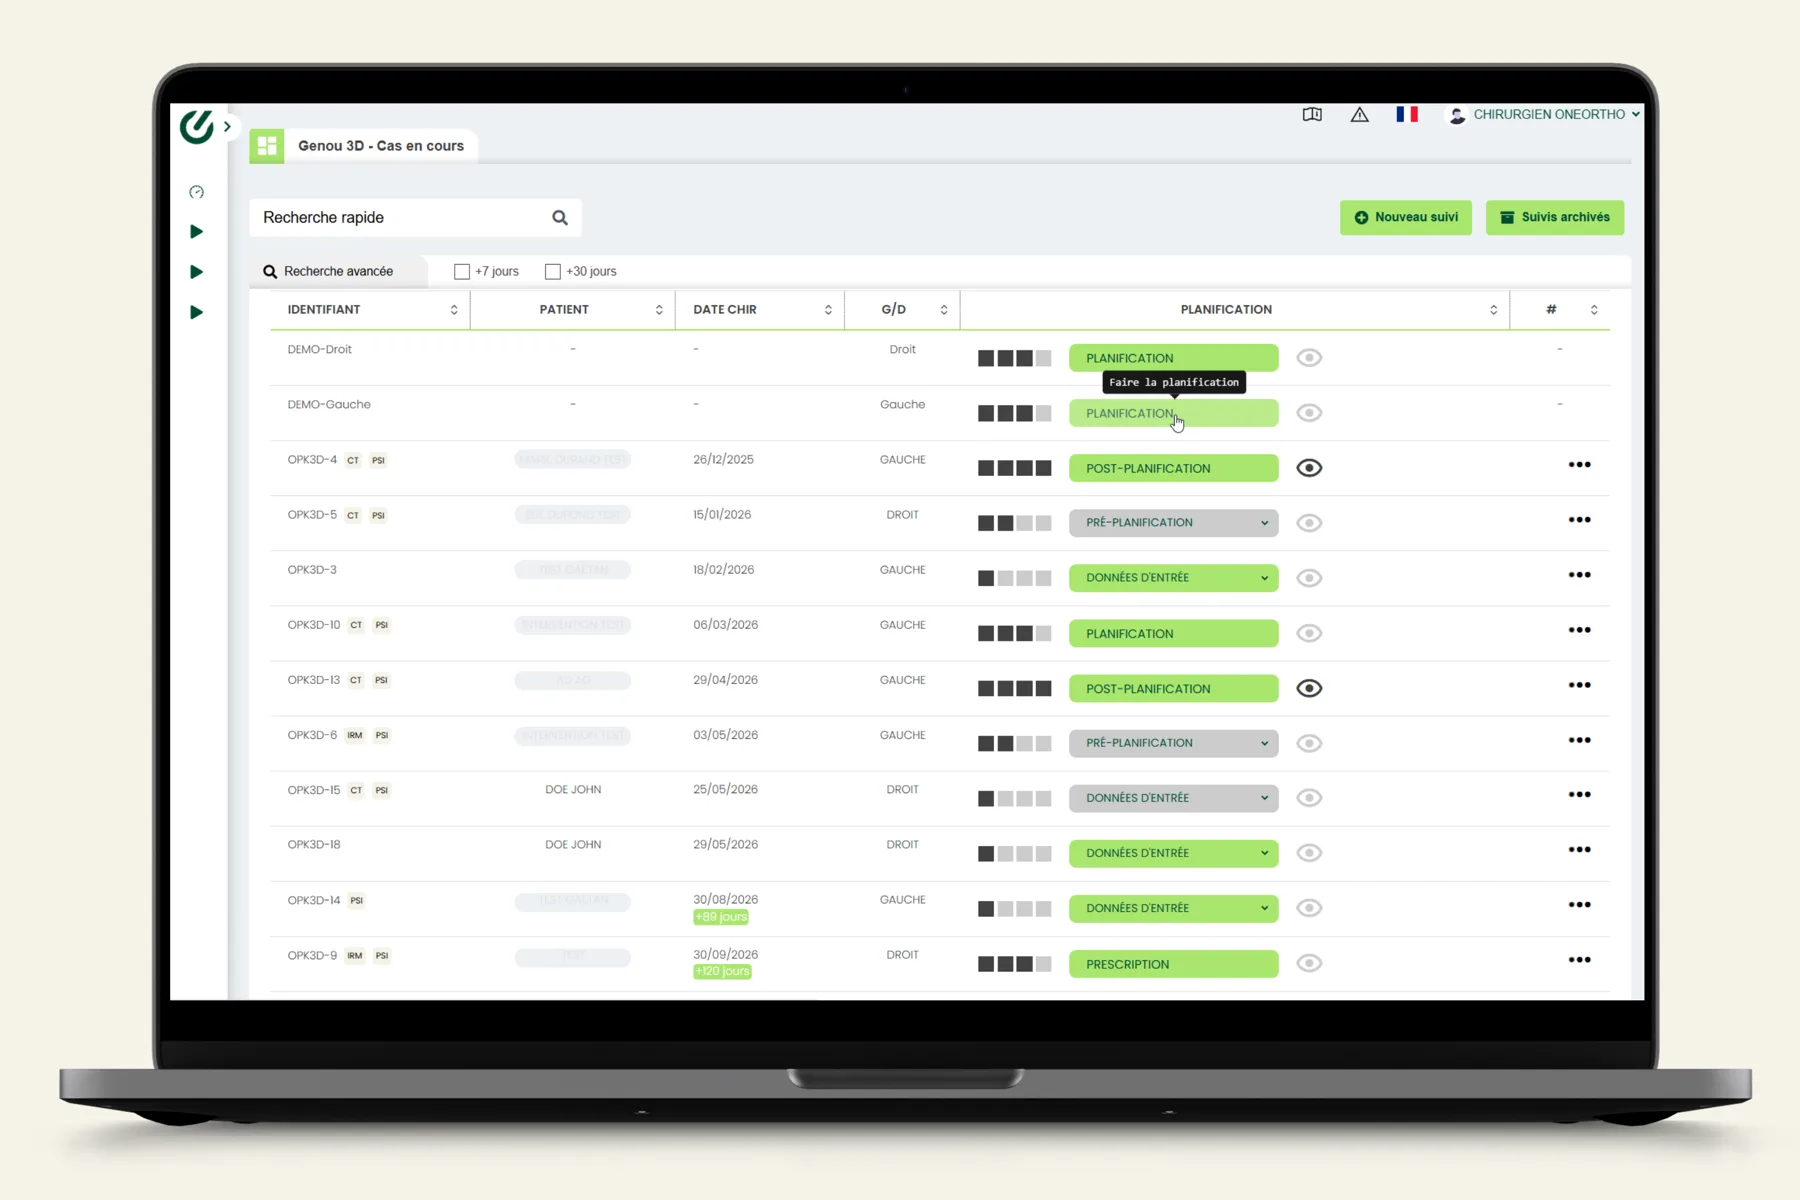
Postoperative data collection via the platform
What this changes for you
Bringing a digital medical device to market: on your own or with OneOrtho Medical's support
None of these steps are optional: they all stem from Regulation (EU) 2017/745 (MDR). You can handle them in-house or entrust them to us.
Without OneOrtho medical
A costly process
Setting up a quality management system and having it certified, within a software scope
Compiling a complete technical file
Contracting with a notified body (NB) and extending your CE scope to software
Organizing a compliant release into production and its control
Setting up vigilance and post-market surveillance
With OneOrtho Medical's support
What we take on
The quality system exists, certified since 2014 and regularly audited
We compile and maintain the technical file
The relationship with the notified body is already in place
Production release and its control are operated on our site
We handle vigilance and post-market follow-up
You bring the technology and the protocol. You keep your intellectual property and your brand.